




Vytas retweeted

We have published the prelim. speaker lineup and program over at the event page, we're incredibly excited for this lineup (including a keynote from Max Welling @wellingmax)! bit.ly/alchemy-2024
This Post is from an account that no longer exists. Learn more
Vytas retweeted

🥳🎉 Registration for the Hünfeld #Workshop "Computer Simulation and #Theory of #Macromolecules" is now open! Don't miss your chance to join us! For more details see 🔗mpinat.mpg.de/workshop/huenf…

Vytas retweeted

Reminder: Our nonprofit, the Open Molecular Software Foundation, is hiring a research software engineer and a technical lead. Please encourage talented science software folks to apply! omsf.io/about/careers/
Registration to the 2024 Workshop on Free Energy Methods in Drug Design is now open:
ssl.eventilla.com/attend/d1L…


The 2024 edition of the Free Energy Workshop will take place in Leiden, Netherlands, 13-15th May, 2024!
We have not yet opened registration, but you can already book the dates in your calendars.
Keep an eye on the wiki for further information: alchemistry.org/wiki/2024_Wo…
#alchemy2024
The 2024 edition of the Free Energy Workshop will take place in Leiden, Netherlands, 13-15th May, 2024!
We have not yet opened registration, but you can already book the dates in your calendars.
Keep an eye on the wiki for further information: alchemistry.org/wiki/2024_Wo…
#alchemy2024
Introducing the organizing committee for #alchemy2024:
Barbara Zarzycka, Bert de Groot, David Hahn, Dima Lupyan, @HighSpeedMode, @hugogdtc, @jenkescheen, @VytasGapsys, @Willemjespers
Thanks everyone involved! It was great to work together on this.
What we built here is an automated workflow for free energy calculations: put ligand SMILES in, get dG back.
With this in hand we probed how various docking protocols would affect dG accuracy

.@VytasGapsys and @jhmchem and colleagues develop an end-to-end relative free energy workflow based on a non-equilibrium switching approach that calculates the binding free energies starting from SMILES strings.
nature.com/articles/s42004-0…
Our latest work on alchemical free energies. This time we used them as an oracle in an active learning loop to navigate chemical space in search for high affinity binders.
Thanks to Yuriy, @gtresadern, David and Bert!
pubs.acs.org/doi/10.1021/acs…
Several main findings below

We were also able to control the way in which to explore the chemical libraries. By changing ligand selection strategy we could
a) quickly find a deep local minimum of strong binders or
b) obtain a good overall description of the chemical space
Vytas retweeted

<- Computational Chemist with background in Quantum Chemistry, protein Molecular Dynamics and industrial experience in Material Science searching for a new position. Ideally in the Karlsruhe/Heidelberg area but not limited to it. Find my LinkedIn profile: linkedin.com/in/martin-werne…
Vytas retweeted

Pre-Exascale Computing of Protein–Ligand Binding #FreeEnergies with Open Source Software for #DrugDesign #cheminformatics
pubs.acs.org/doi/10.1021/acs…
@davidlmobley @gtresadern
#current_issue #JCIM #Letters

Here is something unexpected: just discovered several hidden rules for the @JCIM_JCTC table of contents figure. Can you guess what two rules are violated by this TOC image?

Our latest study on ligand alchemy now in @JCIM_ACS with David @gtresadern @davidlmobley Markus and Bert:
pubs.acs.org/doi/10.1021/acs…
The question we asked here: how far can we push the limits of scaling such calculations relying on the open source software and force fields?

It was great to see that the alchemical protocols parallelize really efficiently: we could make use of ~500 nodes of the Max Planck new supercomputer 'Raven' for 3 days (more than 3 million core hours).
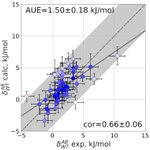
As a result, we also made free energy predictions for the systems in @Ch_Schindler's dataset with the open source force fields, including @openforcefield
